World supplier of antibodies, antigens and kits since 2007

Cell Signal Transduction Research Antibodies
HEXA Polyclonal Antibody
Polyclonal, Rabbit
$225/ 100ul
QuantityTotal: $225
Ship worldwide. Contact order@enogene.com for ordering and bulk pricing or contact your local distributor.
Product Information
Molecular Weight58190Da
Protein NameBeta-hexosaminidase subunit alpha
Human Gene Id3073
Human Swiss Prot NoP06865
Mouse Gene Id15211
Mouse Swiss Prot NoP29416
Rat Gene Id300757
Rat Swiss Prot NoQ641X3
SpecificityHEXA Polyclonal Antibody detects endogenous levels of HEXA protein.
FormulationLiquid in PBS containing 50% glycerol, 0.5% BSA and 0.02% sodium azide.
Concentration1 mg/ml
Storage Stability-20°C/1 year
Other NameHEXA; Beta-hexosaminidase subunit alpha; Beta-N-acetylhexosaminidase subunit alpha; Hexosaminidase subunit A; N-acetyl-beta-glucosaminidase subunit alpha
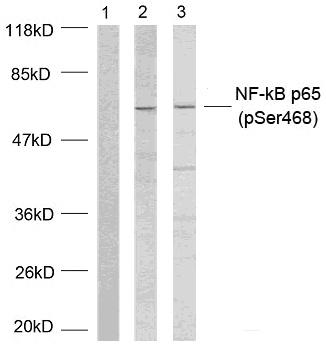
Observed Band(KD)60
Functioncatalytic activity:Hydrolysis of terminal non-reducing N-acetyl-D-hexosamine residues in N-acetyl-beta-D-hexosaminides.,disease:Defects in HEXA are the cause of GM2-gangliosidosis type 1 (GM2G1) [MIM:272800]; also known as Tay-Sachs disease. GM2-gangliosidosis is an autosomal recessive lysosomal storage disease marked by the accumulation of GM2 gangliosides in the neuronal cells. GM2G1 is characterized by GM2 gangliosides accumulation in the absence of HEXA activity, leading to neurodegeneration and, in the infantile form, death in early childhood. GM2G1 has an increased incidence among Ashkenazi Jews and French Canadians in eastern Quebec. It exists in several forms: infantile (most common and most severe), juvenile and adult (late onset).,function:Responsible for the degradation of GM2 gangliosides, and a variety of other molecules containing terminal N-acetyl hexosamines, in the brain
Subcellular LocationLysosome.
ExpressionBrain,Eye,Liver,Ovary,Uterus,
PurificationThe antibody was affinity-purified from rabbit antiserum by affinity-chromatography using epitope-specific immunogen.
ImmunogenSynthesized peptide derived from HEXA . at AA range: 121-170
Gene NameHEXA
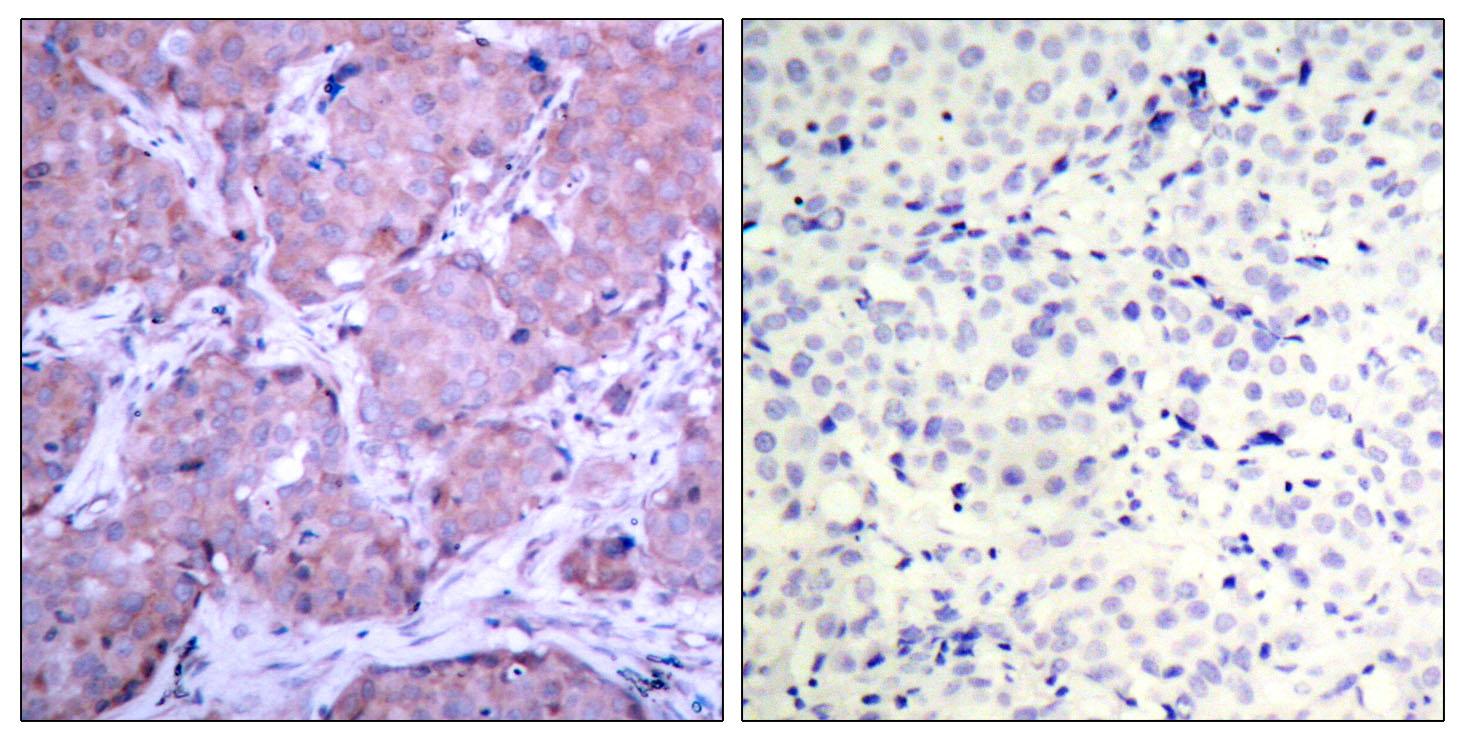
DilutionWestern Blot: 1/500 - 1/2000. IHC-p: 1:100-300 ELISA: 1/20000. Not yet tested in other applications.
Backgroundhexosaminidase subunit alpha(HEXA) Homo sapiens This gene encodes a member of the glycosyl hydrolase 20 family of proteins. The encoded preproprotein is proteolytically processed to generate the alpha subunit of the lysosomal enzyme beta-hexosaminidase. This enzyme, together with the cofactor GM2 activator protein, catalyzes the degradation of the ganglioside GM2, and other molecules containing terminal N-acetyl hexosamines. Mutations in this gene lead to an accumulation of GM2 ganglioside in neurons, the underlying cause of neurodegenerative disorders termed the GM2 gangliosidoses, including Tay-Sachs disease (GM2-gangliosidosis type I). Alternative splicing results in multiple transcript variants, at least one of which encodes a preproprotein that is proteolytically processed. [provided by RefSeq, Jan 2016],


Conjugated Antibody
This product is also available conjugated. Format: E20-75233-Conjugate. Price: $485 / Size: 100μl.
E20-75233-FITC$485 / 100μlE20-75233-Biotin$485 / 100μlE20-75233-HRP$485 / 100μlE20-75233-AP$485 / 100μlE20-75233-AF350$485 / 100μlE20-75233-AF405M$485 / 100μlE20-75233-AF488$485 / 100μlE20-75233-AF514$485 / 100μlE20-75233-AF532$485 / 100μlE20-75233-AF555$485 / 100μlE20-75233-AF568$485 / 100μlE20-75233-AF405S$485 / 100μlE20-75233-AF405L$485 / 100μlE20-75233-AF546$485 / 100μlE20-75233-AF594$485 / 100μlE20-75233-AF610$485 / 100μlE20-75233-AF635$485 / 100μlE20-75233-AF647$485 / 100μlE20-75233-AF680$485 / 100μlE20-75233-AF700$485 / 100μlE20-75233-AF750$485 / 100μlE20-75233-AF790$485 / 100μlE20-75233-APC$485 / 100μlE20-75233-PE$485 / 100μlE20-75233-Cy3$485 / 100μlE20-75233-Cy5$485 / 100μlE20-75233-Cy5.5$485 / 100μlE20-75233-Cy7$485 / 100μlE20-75233-PE-Cy5$485 / 100μl
Contact order@enogene.com to order conjugated formats. Magnetic Beads (1-30μm) conjugation is also available.


Categories by research area
Categories by product type
- Antibodies
- Tag antibodies
- Internal control antibodies
- Pathological (Immunohistochemistry, IHC) antibodies
- Cell Signal Transduction Research Antibodies
- Secondary Antibody
- Antibody Sampler Kits
- Recombinant Proteins
- Fluorescence Reagents
- Signal Inhibitors and Activators
- Transfection Reagent
- Cell biology kits