
You are here: EnoGene Home > Products > Cell Signal Transduction Research Antibodies > OCRL Polyclonal Antibody
OCRL Polyclonal Antibody
Datasheet
Catalog Number:
E20-73228Amount:
100ulMolecular Weight:
104205DaPurification:
The antibody was affinity-purified from rabbit antiserum by affinity-chromatography using epitope-specific immunogen.Immunogen:
The antiserum was produced against synthesized peptide derived from human OCRL. AA range:150-199Source:
Polyclonal, Rabbit
- Western Blot analysis of various cells using OCRL Polyclonal Antibody
Gene Name:
OCRLDilution:
Western Blot: 1/500 - 1/2000. ELISA: 1/10000. Not yet tested in other applications.Protein Name:
Inositol polyphosphate 5-phosphatase OCRL-1Human Gene Id:
4952Human Swiss Prot No:
Q01968Specificity:
OCRL Polyclonal Antibody detects endogenous levels of OCRL protein.Formulation:
Liquid in PBS containing 50% glycerol, 0.5% BSA and 0.02% sodium azide.Concentration:
1 mg/mlStorage Stability:
-20°C/1 yearOther Name:
OCRL; INPP5F; OCRL1; Inositol polyphosphate 5-phosphatase OCRL-1; Lowe oculocerebrorenal syndrome proteinObserved Band(KD):
104Background:
OCRL, inositol polyphosphate-5-phosphatase(OCRL) Homo sapiens This gene encodes an inositol polyphosphate 5-phosphatase. This protein is involved in regulating membrane trafficking and is located in numerous subcellular locations including the trans-Golgi network, clathrin-coated vesicles and, endosomes and the plasma membrane. This protein may also play a role in primary cilium formation. Mutations in this gene cause oculocerebrorenal syndrome of Lowe and also Dent disease. Alternate splicing results in multiple transcript variants. [provided by RefSeq, Jan 2016],Function:
catalytic activity:1-phosphatidyl-1D-myo-inositol 4,5-bisphosphate + H(2)O = 1-phosphatidyl-1D-myo-inositol 4-phosphate + phosphate.,caution:It is uncertain whether Met-1, Met-18 or Met-20 is the initiator.,disease:Defects in OCRL are the cause of Dent disease type 2 (DD2) [MIM:300555]. DD2 is a renal disease belonging to the 'Dent disease complex', a group of disorders characterized by proximal renal tubular defect, hypercalciuria, nephrocalcinosis, and renal insufficiency. The spectrum of phenotypic features is remarkably similar in the various disorders, except for differences in the severity of bone deformities and renal impairment. Characteristic abnormalities include low-molecular-weight proteinuria and other features of Fanconi syndrome, such as glycosuria, aminoaciduria, and phosphaturia, but typically do not include proximal renal tubular acidosis. Progressive renal failure is cSubcellular Location:
Cytoplasmic vesicle, phagosome membrane . Early endosome membrane . Membrane, clathrin-coated pit . Cell projection, cilium, photoreceptor outer segment . Cell projection, cilium . Cytoplasmic vesicle . Endosome . Golgi apparatus, trans-Golgi network . Lysosome . Also found on macropinosomes (PubMed:25869668). Colocalized with APPL1 on phagosomes (PubMed:22072788). .Expression:
Brain, skeletal muscle, heart, kidney, lung, placenta and fibroblasts. Expressed in the retina and the retinal pigment epithelium.

Related Products
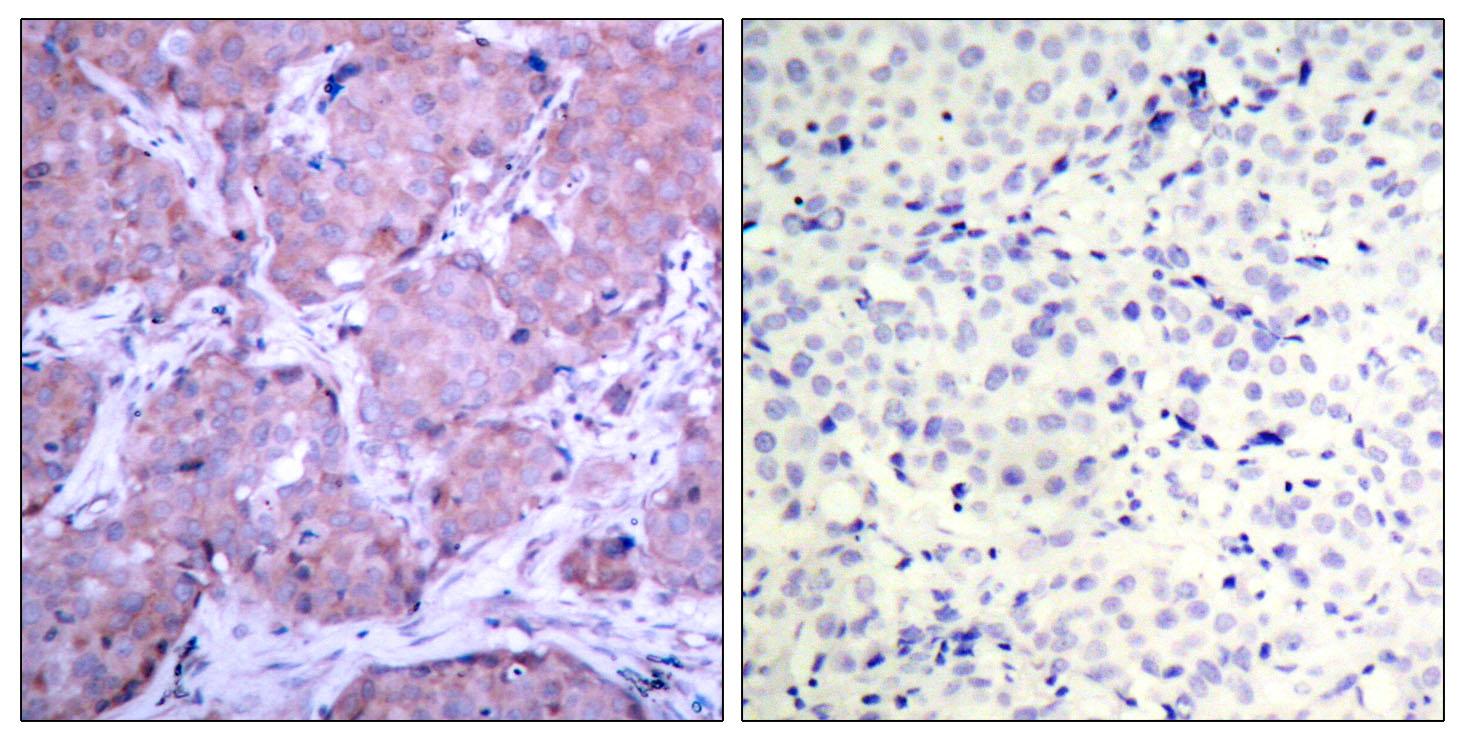
- PTEN (Phospho-Ser380) Antibody Datasheet
Catalog Number:
E011009-1Amount:
50μg/50μlReactivity:
Human, Mouse, RatApplications:
IHC: 1:50~1:100Swiss-Prot No.:
P60484Price:
145$
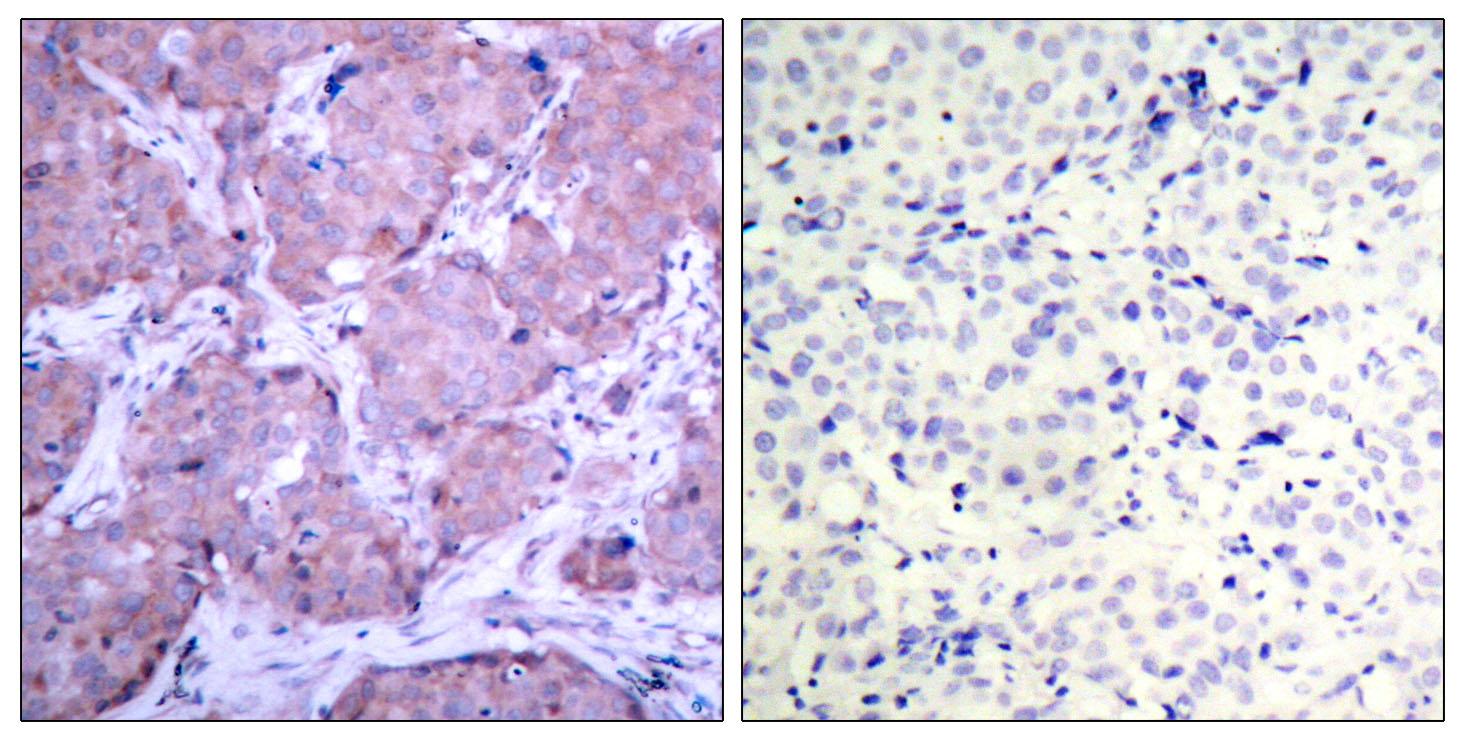
- PTEN (Phospho-Ser380) Antibody Datasheet
Catalog Number:
E011009-2Amount:
100μg/100μlReactivity:
Human, Mouse, RatApplications:
IHC: 1:50~1:100Swiss-Prot No.:
P60484Price:
225$
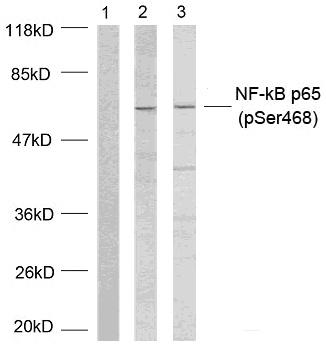
- NFκB-p65 (Phospho-Ser468) Antibody Datasheet
Catalog Number:
E011013-1Amount:
50μg/50μlReactivity:
Human, Mouse, RatApplications:
WB: 1:500~1:1000 IF:1:100~1:200Swiss-Prot No.:
Q04206Price:
145$
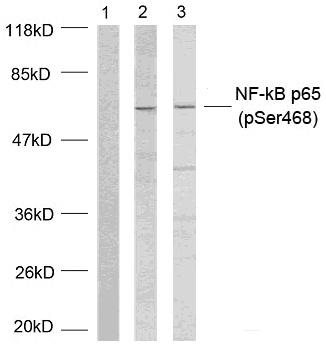
- NFκB-p65 (Phospho-Ser468) Antibody Datasheet
Catalog Number:
E011013-2Amount:
100μg/100μlReactivity:
Human, Mouse, RatApplications:
WB: 1:500~1:1000 IF:1:100~1:200Swiss-Prot No.:
Q04206Price:
225$

- Rel (Phospho-Ser503) Antibody Datasheet
Catalog Number:
E011020-1Amount:
50μg/50μlReactivity:
HumanApplications:
IF:1:100~1:200Swiss-Prot No.:
Q04864Price:
145$

Catalog Number
- Amount:100ul
- Price:225$

Contact us
Categories by research area
Categories by product type
- Antibodies
- Tag antibodies
- Internal control antibodies
- Pathological (Immunohistochemistry, IHC) antibodies
- Cell Signal Transduction Research Antibodies
- Secondary Antibody
- Antibody Sampler Kits
- Recombinant Proteins
- Fluorescence Reagents
- Signal Inhibitors and Activators
- Transfection Reagent
- Cell biology kits