World supplier of antibodies, antigens and kits since 2007

Other Products
DMGDH Polyclonal Antibody
Human;Rat;Mouse;
WB;IHC-p;IF(paraffin section);ELISA
Polyclonal, Rabbit,IgG
Price on request
Quantity
Ship worldwide. Contact order@enogene.com for ordering and bulk pricing or contact your local distributor.
Gene NameDMGDH
Product Information
Protein NameDimethylglycine dehydrogenase mitochondrial
Human Gene Id29958
Human Swiss Prot NoQ9UI17
Mouse Swiss Prot NoQ9DBT9
SpecificityDMGDH Polyclonal Antibody detects endogenous levels of DMGDH protein.
FormulationLiquid in PBS containing 50% glycerol, 0.5% BSA and 0.02% sodium azide.
Concentration1 mg/ml
Storage Stability-20°C/1 year
Other NameDMGDH; Dimethylglycine dehydrogenase; mitochondrial; ME2GLYDH
Molecular Weight96807Da
Observed Band(KD)97
Functioncatalytic activity:N,N-dimethylglycine + acceptor + H(2)O = sarcosine + formaldehyde + reduced acceptor.,cofactor:Binds 1 FAD covalently per monomer.,disease:Defects in DMGDH are the cause of DMGDH deficiency (DMGDHD) [MIM:605850]. DMGDHD is a disorder characterized by fish odor, muscle fatigue with increased serum creatine kinase. Biochemically it is characterized by an increase of N,N-dimethylglycine (DMG) in serum and urine.,pathway:Amine and polyamine degradation; betaine degradation; sarcosine from betaine: step 2/2.,similarity:Belongs to the gcvT family.,subunit:Monomer.,
Subcellular LocationMitochondrion.
ExpressionKidney,Trachea,
ImmunogenThe antiserum was produced against synthesized peptide derived from human DMGDH. AA range:817-866
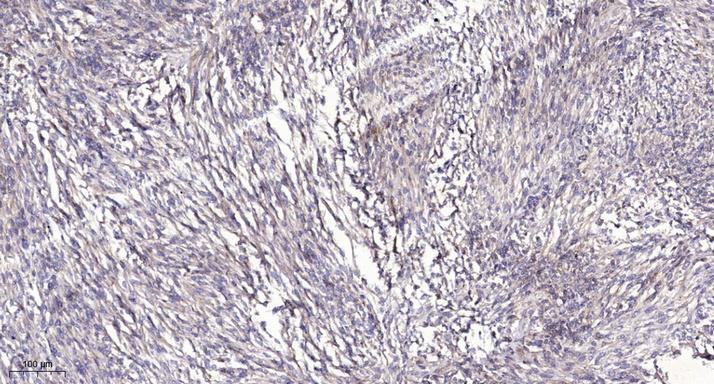
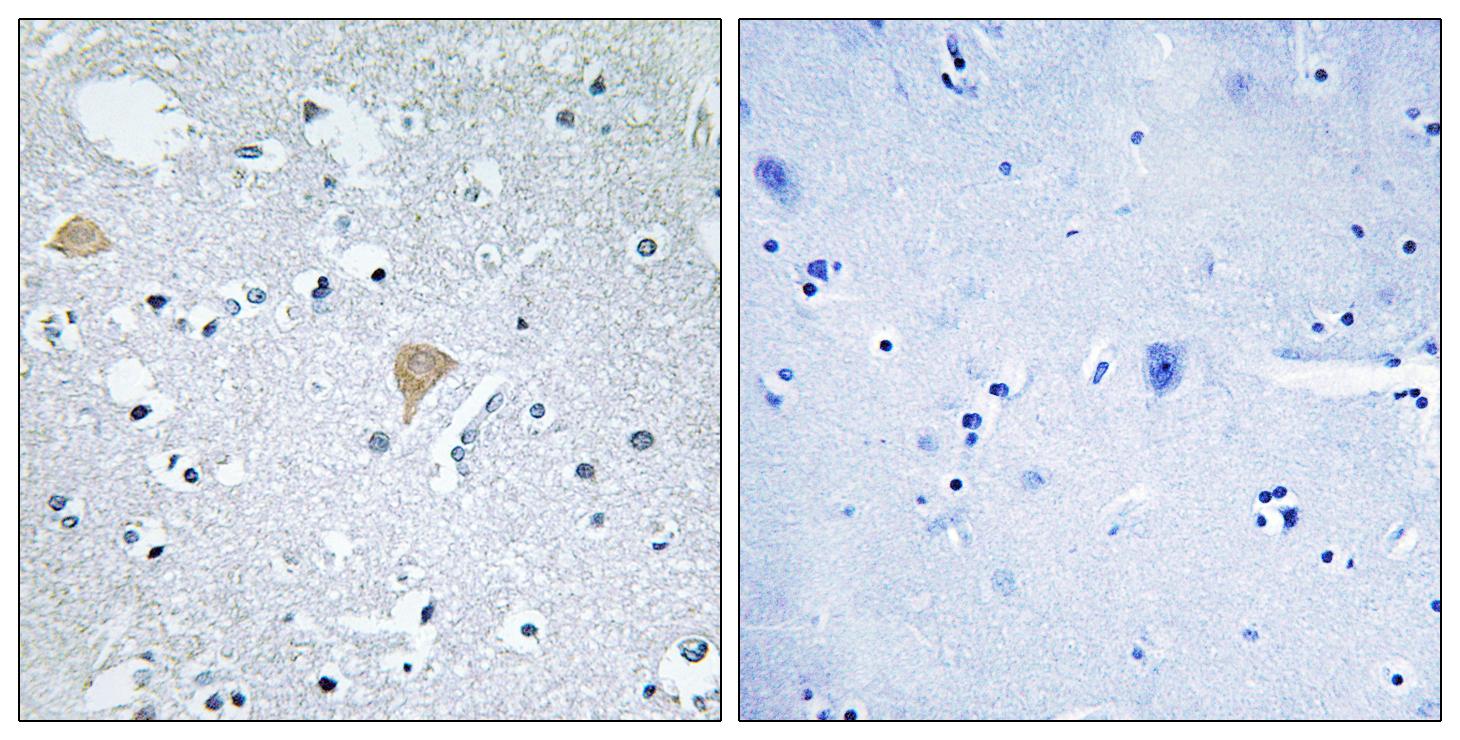
DilutionWestern Blot: 1/500 - 1/2000. Immunohistochemistry: 1/100 - 1/300. ELISA: 1/20000. Not yet tested in other applications.
PurificationThe antibody was affinity-purified from rabbit antiserum by affinity-chromatography using epitope-specific immunogen.
BackgroundThis gene encodes an enzyme involved in the catabolism of choline, catalyzing the oxidative demethylation of dimethylglycine to form sarcosine. The enzyme is found as a monomer in the mitochondrial matrix, and uses flavin adenine dinucleotide and folate as cofactors. Mutation in this gene causes dimethylglycine dehydrogenase deficiency, characterized by a fishlike body odor, chronic muscle fatigue, and elevated levels of the muscle form of creatine kinase in serum. Alternative splicing results in multiple transcript variants. [provided by RefSeq, Jul 2013],
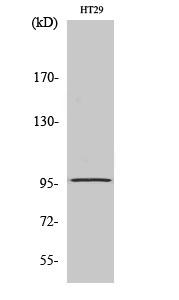
Western Blot analysis of various cancer. 1, cells using DMGDH Polyclonal pH9.0 was Antibody. diluted at 1:200
Conjugated Antibody
This product is also available conjugated. Format: E20-713862-Conjugate. Price: $485 / Size: 100μl.
E20-713862-FITC$485 / 100μlE20-713862-Biotin$485 / 100μlE20-713862-HRP$485 / 100μlE20-713862-AP$485 / 100μlE20-713862-AF350$485 / 100μlE20-713862-AF405M$485 / 100μlE20-713862-AF488$485 / 100μlE20-713862-AF514$485 / 100μlE20-713862-AF532$485 / 100μlE20-713862-AF555$485 / 100μlE20-713862-AF568$485 / 100μlE20-713862-AF405S$485 / 100μlE20-713862-AF405L$485 / 100μlE20-713862-AF546$485 / 100μlE20-713862-AF594$485 / 100μlE20-713862-AF610$485 / 100μlE20-713862-AF635$485 / 100μlE20-713862-AF647$485 / 100μlE20-713862-AF680$485 / 100μlE20-713862-AF700$485 / 100μlE20-713862-AF750$485 / 100μlE20-713862-AF790$485 / 100μlE20-713862-APC$485 / 100μlE20-713862-PE$485 / 100μlE20-713862-Cy3$485 / 100μlE20-713862-Cy5$485 / 100μlE20-713862-Cy5.5$485 / 100μlE20-713862-Cy7$485 / 100μlE20-713862-PE-Cy5$485 / 100μl
Contact order@enogene.com to order conjugated formats. Magnetic Beads (1-30μm) conjugation is also available.


Categories by research area
Categories by product type
- Antibodies
- Primary Antibodies
- Tag antibodies
- Internal control antibodies
- Pathological (Immunohistochemistry, IHC) antibodies
- Cell Signal Transduction Research Antibodies
- Secondary Antibody
- Antibody Sampler Kits
- ELISA Kits
- Recombinant Proteins
- Fluorescence Reagents
- Signal Inhibitors and Activators
- Transfection Reagent
- Cell biology kits
- Overexpression Cell Lines
- Cell Lysates
- Purification Resins
- 2019-nCoV & Monkeypox